A versatile resource for normalizing RNA-sequencing data
A critical step when analyzing RNA-seq data is normalization. Normalization removes systematic biases that arise from variability in experimental conditions, sample collection and preparation, and machine parameters, etc., while preserving the variation in gene expression that occurs because of biologically relevant changes in transcription at the same time.
RNA-seq Normalization implements seven normalization methods, among which five are existing methods (DESeq, TMM, RPM, PoissonSeq and UQ) for fresh frozen (FF) RNA-seq data. Two novel normalization methods, MIXnorm and SMIXnorm, can be applied to both FF and formalin-fixed paraffin-embedded (FFPE) RNA-seq data.
Architect of Our Tools
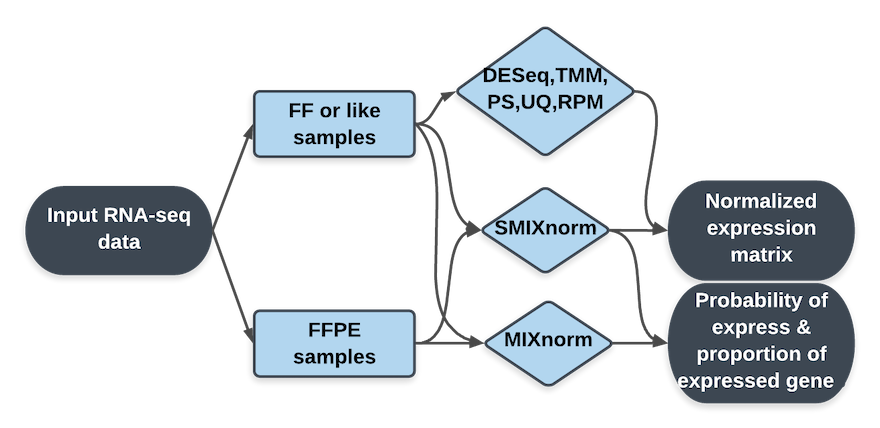
All methods take raw count matrix as input and reture the normalized expression matrix as a downloadable csv file. Offline versions of MIXnorm and SMIXnom are available from the Download page. The probability of being expressed for each gene and the proportion of expressed genes are also returned from the offline version.